
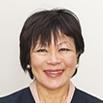
Swee Lay Thein, B.S., F.R.C.P., F.R.C.Path., D.Sc., FMedSci
Senior Investigator
Sickle Cell Genetics and Pathophysiology
NHLBI
NIH Distinguished Clinical Investigator
NHLBI
Research Topics
The Thein group conducts research on the pathophysiology of sickle cell disease (SCD) applying insights from genotype-phenotype observations to accelerate translation of basic discovery to therapeutic applications. By combining clinical, laboratory, imaging and new diagnostic techniques, we hope to improve phenotypic definition of SCD at all levels and to introduce more targeted pharmaceutical therapies for SCD. The Thein laboratory is also involved in pre-clinical and clinical testing of drugs and innovative compounds that reduce sickle hemoglobin (HbS) polymerization, improve red cell rheology, or induce fetal hemoglobin.
SCD is an inherited hemoglobin disorder prevalent in regions where malaria was historically endemic, including sub-Saharan Africa, India, the Middle East, and the Mediterranean. Currently, an estimated 300,000 affected babies are born each year, more than 80% of whom are in Africa. In the United States, SCD is considered “rare”, but nonetheless affects an estimated 100,000 Americans (the majority being of African descent) and 1 in 365 African-American births.
The single nucleotide change underpinning SCD belies the extreme variability and severity of SCD complications, that becomes more apparent with aging, and presents major challenges in clinical management. Both acquired and inherited factors contribute to this clinical complexity of SCD. Numerous genetic association studies (candidate gene, genome-wide association (GWA), whole exome and whole genome sequencing) have attempted to identify genetic variants with particular disease complications. In a GWA study in 2007, the Thein group identified the BCL11A gene as a key repressor of gamma-globin gene, a finding that resuscitated the field in fetal hemoglobin regulation and therapeutic re-activation of gamma-globin genes for the beta-hemoglobinopathies. The finding has led to genetic strategies targeting BCL11A regulation for therapeutic reactivation in SCD and beta thalassemia under clinical development with promising results.
Prediction of disease severity and clinical course of SCD has been the topic of many reviews and, to date there is no clear algorithm using genetic and/or imaging, and/or laboratory markers that can reliably predict mortality risk in SCD. The clinical implications of delineating the genetic modifiers of SCD are significant; it will provide us the ability to predict disease severity based on a genetic SCD “panel”, and dissecting the role of new genetic modifiers might suggest new therapeutic targets for investigation. A primary approach for disease risk prediction has been polygenic risk scoring but the results remain moderately successful; they may not be so applicable in a complex disorder such as SCD that has numerous potential predictor variables. Towards the advancement of personalized medicine, the Thein group is exploring new machine learning methods incorporating predictive biomarkers (laboratory, imaging, and physiological) with genetic (germline and somatic) risk variants into risk assessment algorithms to improve risk stratification before end organ damage sets in.
While we apply insights from the basic research to identify new therapeutic targets, the Thein lab is also involved in a project with Dr Bill Eaton (Laboratory of Chemical Physics, NIDDK/NIH) to discover newer anti-sickling agents using high throughput screening of drug libraries. Not only will the compounds provide further insight on the sickling mechanisms, but those that show therapeutically significant effects at concentrations known to be non-toxic can be very rapidly approved for clinical trials as these agents are either already FDA-approved or under clinical development for other diseases. It could well be that this small molecule game changer for SCD already exists among the drugs that are being used for other disorders. Furthermore, unravelling the mechanisms of the anti-sickling effects of these agents could well reveal fresh insights on the HbS fiber formation.
Biography
Swee Lay Thein was educated in both Malaysia and the United Kingdom. She completed her specialist training in hematology at the U.K. Royal Postgraduate Medical School, Hammersmith, and the Royal Free Hospital, London. In 1982, she joined the U.K. Medical Research Council Molecular Hematology Unit in Oxford, rising from MRC clinical training fellow, Wellcome Senior Fellow in Clinical Science, to MRC senior clinical scientist, and honorary consultant hematologist.
In 2000, Dr Thein became professor of molecular hematology and consultant hematologist at King’s College London / King’s College Hospital, where she served as clinical director of the Red Cell Centre, treating adults with sickle cell disease and advising clinicians and researchers worldwide on unusual forms of thalassemias and inherited anemias.
Dr. Thein joined the NHLBI in spring 2015 as Senior Investigator and Chief of the institute’s newly formed Sickle Cell Branch, later consolidated into the Hematology branch in summer 2026.
Dr. Thein has published over 400 scientific articles and supervised numerous graduate students and fellows. Since 2006, she has directed and hosted the Sickle Cell in Focus Conference, an annual 2-day international conference. She has served on editorial boards, national and international advisory committees, and has been recognized with major awards. Her honors include Fellowship of the Academy of Medical Sciences (FMedSci, 2003), UK; recognition of her research by scientists as one of the top 10 science stories (The Guardian, 2022), co-recipient of the Shaw Prize in Life Sciences & Medicine (2024); the Helen M. Ranney award upon her induction into the Association of American Physicians (2025); election to the Fellowship of the Royal Society (2025); co-recipient of the Breakthrough Prize in Life Sciences and the Warren Alpert Foundation Prize in 2026.
Her research focuses on how genetic mutations and modifiers affect sickle cell disease severity, using genomic and pathophysiologic approaches to improve diagnosis and treatment, with the goal of translating discoveries into new therapies beyond hydroxyurea. Dr. Thein’s leadership and scientific innovations have fundamentally advanced the understanding and management of hemoglobinopathies worldwide.
Selected Publications
- Conrey A, Asomaning N, Frey I, Charles RP, Lovins D, Xu JZ, Mendez-Marti S, Le K, Kruah B, Li Q, Dunkelberger E, Cellmer T, Yates A, Wind-Rotolo M, Huston C, Jeffries N, Eaton WA, Thein SL. Long-term mitapivat treatment is safe and efficacious in patients with sickle cell disease. Blood Red Cells Iron. 2025;1(2).
- D'Alessandro A, Le K, Lundt M, Li Q, Dunkelberger EB, Cellmer T, Worth AJ, Patil S, Huston C, Grier A, Dzieciatkowska M, Stephenson D, Eaton WA, Thein SL. Functional and multi-omics signatures of mitapivat efficacy upon activation of pyruvate kinase in red blood cells from patients with sickle cell disease. Haematologica. 2024;109(8):2639-2652.
- Le K, Wang X, Chu J, Lundt M, Lee YY, Conrey A, Frey I, Giannini S, Kosinski PA, Hausman JM, Low PS, Jeffries N, Desai SA, Thein SL. Activating pyruvate kinase improves red blood cell integrity by reducing band 3 tyrosine phosphorylation. Blood Adv. 2024;8(21):5653-5662.
- Ray R, Li H, Gao S, Asomaning N, Le K, Ahmad MM, Wang X, Li Y, Kamimura S, Dalgard CL, Liu C, Quezado ZMN, Lack J, Tumburu L, Thein SL. Variations in mitochondrial genome as potential prognostic markers in sickle cell disease. Haematologica. 2026.
- Parekh DS, Eaton WA, Thein SL. Recent developments in the use of pyruvate kinase activators as a new approach for treating sickle cell disease. Blood. 2024;143(10):866-871.
Related Scientific Focus Areas
Molecular Biology and Biochemistry
View additional Principal Investigators in Molecular Biology and Biochemistry
This page was last updated on Thursday, October 8, 2026